We use cookies to run this site and, with your consent, to understand how it is used and to follow up with companies interested in our work. You can change your choice at any time under “Cookie settings”. Cookie Policy · Privacy Policy · Imprint
Streamline pharma workflows, from batch records to safety reports.
Batch reviews, regulatory submissions and safety reporting depend on complete, consistent documentation. Teams still spend hours gathering records, checking data and preparing files across systems.
Our AI agents automate that work end-to-end within your RIM, QMS, safety database and ERP, flagging gaps and preparing documentation for expert review. Release decisions and clinical assessments stay with qualified professionals.
File & maintainDossiers assembled, variations classified, renewals tracked in every market1
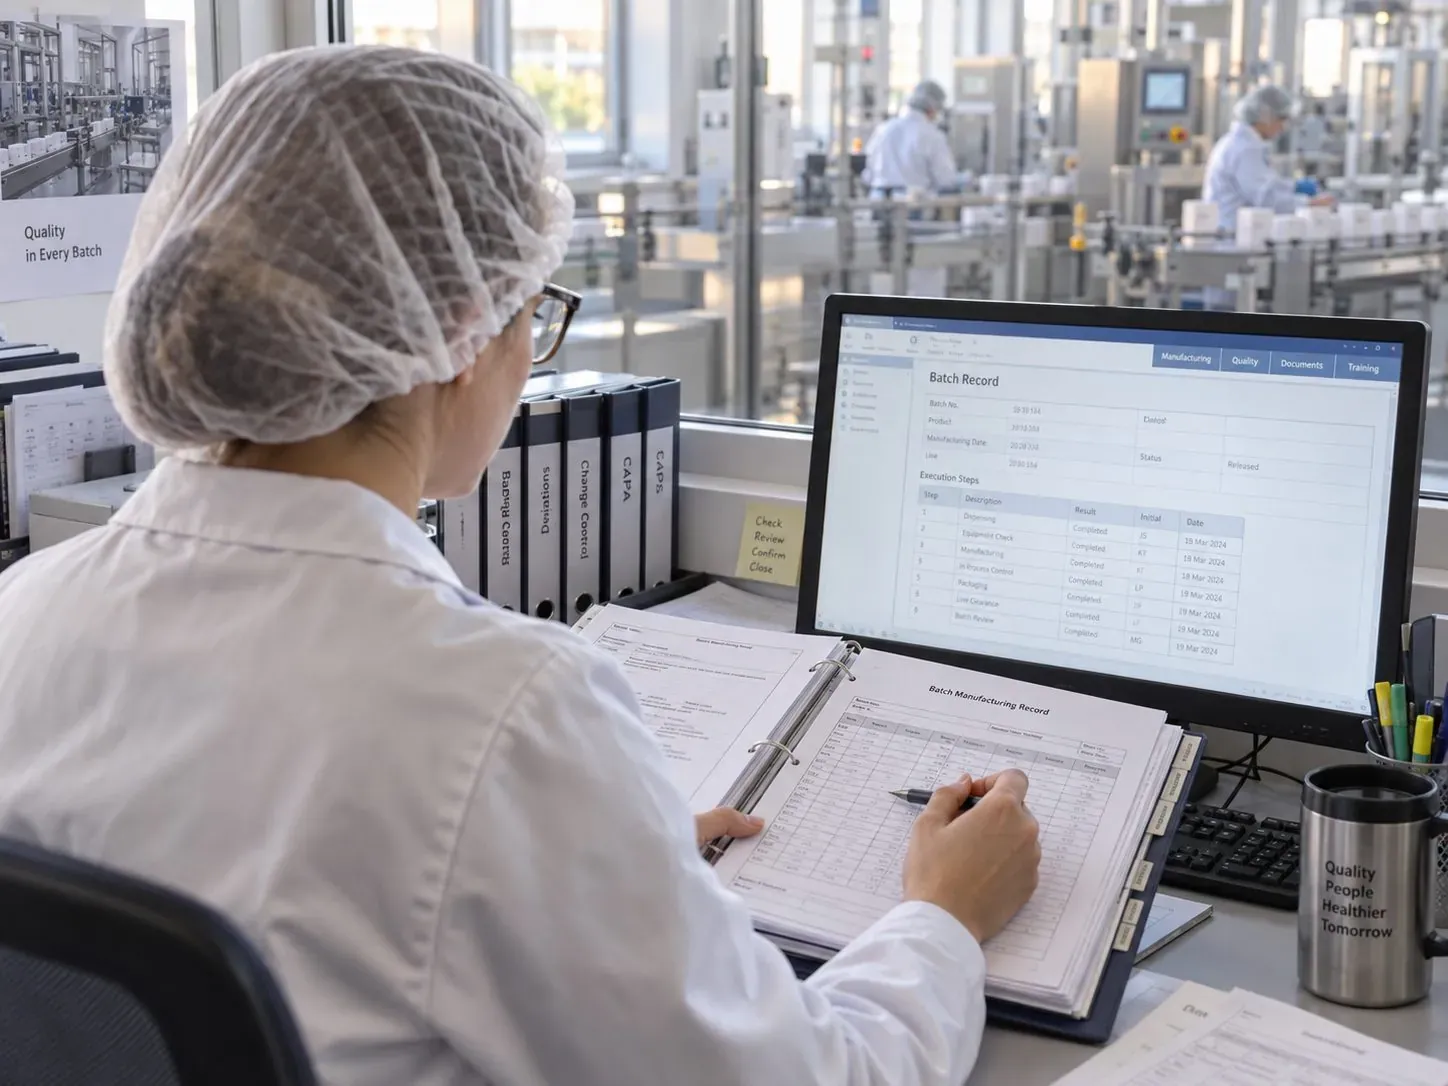
Review & releaseBatch records checked, deviations investigated, CAPA tracked to closure2
Report & answerSafety cases coded and submitted, medical inquiries answered from approved content3
WHAT WE DO FOR PHARMACEUTICALS
Faster batch release,
regulatory filings on time.
Agents assemble each submission and report. Your experts decide.
1 / 3
Production volume · EU pharma vs all manufacturing, 2022 to 2025Pharma output up 28%, manufacturing down 2%. Every batch adds records to review.
Batch records reviewed before the Qualified Person signs.
AI agents check each executed batch record against the master, draft the deviation investigation and track CAPA. Your Qualified Person decides on a short list.
Batch record review · Deviations · CAPA
Post-authorization variation scopes started · EU centrally authorized medicines, 20253,831 Type IA, 3,490 Type IB and 1,284 Type II, each classified differently.
3,831Type IA
3,490Type IB
1,284Type II
Regulatory Affairs · automation potential
70–80%
Pharmacovigilance & Safety · automation potential
70–80%
Source: EMA, Monthly statistics report, December 2025
Variations classified, filed and tracked in every market.
Each change is classified, assembled into the submission sequence and tracked against its deadline in every market. AI agents prepare the work, and your regulatory team reviews and files.
CTD assembly · Variations · Case processing
Firms using AI · EU, 202541% of pharma firms use AI, twice the average. The question is which processes, with what validation.
20%All enterprises
41%Pharmaceuticals
Fastest go-live *
from 12 wks
Payback
< 12 mo
Less manual work
5–10×
Source: Eurostat, isoc_eb_ain2 (C21), 2025
Safety, inquiries and serialization, handled under GxP.
Safety reports are triaged, medical inquiries answered from approved responses and serialization data uploaded, each step validated under Annex 11. AI agents keep a full audit trail, and a person signs off.
Safety intake · Medical information · Serialization
* Our fastest go-live. Timelines depend on the complexity of the project and its processes.
INSIDE A PHARMACEUTICAL COMPANY
Custom agents, built for every
pharmaceutical department.
We don't deploy generic software. We design AI Agents around the processes, tools and people of each function, then connect them where it matters. Click any department to explore.
Regulatory Affairs
Dossiers assembled, variations filed, every market current.
Agents collect source documents, build and validate eCTD sequences, classify post-approval changes and prepare variation packages, track renewals and commitments and keep the RIM system current, so regulatory leads spend their time on strategy and agency dialogue.
Agents classify and file eTMF documents and chase gaps, check essential documents before site activation, track enrollment against plan, manage data queries and reconcile serious adverse events between systems, so study teams manage sites instead of paperwork.
Automation potential69–79%
Trial Master File Management
Site Activation & Essential Documents
Patient Recruitment Tracking
Data Query Management
Clinical SAE Reconciliation
Quality Assurance & GMP
Batch records checked, deviations investigated, CAPA closed.
Agents compare executed batch records against the master and flag exceptions, register and draft deviation investigations, track CAPA and effectiveness checks, administer change control and manage controlled documents. The Qualified Person releases; the agent prepares.
Automation potential70–80%
Batch Record Review
Deviation Management
CAPA Tracking & Effectiveness
Change Control Administration
Document Control & SOP Management
Pharmacovigilance & Safety
Cases coded and reported within timelines, signals visible.
Agents capture and triage reports from every source, code events with MedDRA and draft narratives, submit expedited reports to EudraVigilance, screen the literature and compile PSURs, so the QPPV and medical reviewers assess instead of transcribe.
Automation potential70–80%
ICSR Case Intake & Triage
Case Processing & MedDRA Coding
Expedited Reporting to EudraVigilance
Literature Monitoring
PSUR / PBRER Preparation
Medical Affairs & Medical Information
Literature screened, materials reviewed, inquiries answered from approved content.
Agents screen publications, route promotional materials through medical, legal and regulatory review and check claims against labeling, compile MSL insights, and triage and draft answers to HCP and patient inquiries from approved standard responses, forwarding adverse events within timelines. Medical judgment stays with the physician.
Automation potential70–80%
Scientific Literature Surveillance
Medical Review of Promotional Material
Medical Science Liaison Activity Reporting
Medical Inquiry Intake & Triage
Standard Response Drafting
Manufacturing, Supply Chain & Serialization
Demand planned, shortages notified early, every pack uploaded.
Agents build the demand and supply plan, prepare batch documentation, monitor inventory and expiry, handle cold-chain excursions for QA decision, prepare shortage notifications per country, upload pack data to the EU Hub and investigate verification alerts, so supply managers decide allocations instead of chasing status.
Agents run material requirements and purchase orders, coordinate orders, forecasts and documents with contract manufacturers, match invoices to orders and receipts, collect the documents QA needs to qualify suppliers and compute supplier scorecards, so buyers negotiate instead of administer.
Automation potential73–83%
Material Requirements & Purchasing
CDMO Coordination
Invoice 3-Way Matching
Supplier Qualification Document Collection
Supplier Performance Scorecards
Business Development & Licensing
Opportunities screened, data rooms indexed, milestones tracked.
Agents screen in-licensing opportunities against the strategy, index licensing data rooms against the due diligence checklist, coordinate functional due diligence, administer alliance committees and track milestones, royalties and partner reporting obligations, so deal teams negotiate instead of compile.
Automation potential70–80%
Opportunity Screening & Pipeline Intelligence
Data Room Review Support
Due Diligence Coordination
Milestone & Royalty Tracking
Partner Reporting Obligations
Commercial Operations & Market Access
Tenders prepared, dossiers assembled, transfers of value disclosed.
Agents monitor tender portals and assemble bids, process wholesaler and hospital orders, compile pricing and reimbursement dossiers, monitor international reference prices, record HCP interactions against compliance rules and compile EFPIA disclosure reports, so commercial and access teams argue the value instead of formatting it.
Automation potential72–82%
Tender Monitoring & Preparation
Customer Order Management
Pricing & Reimbursement Dossier Preparation
International Reference Pricing Monitoring
Transparency & Disclosure Reporting
Finance & Controlling
The close on time, rebates accrued, costs tracked.
Agents process supplier and CDMO invoices, run the close checklist and reconciliations, compute rebate and clawback accruals, track R&D project costs and build the cash forecast, so controllers explain the numbers instead of assembling them.
GMP training assigned, qualifications valid, onboarding done.
Agents assign role-based GMP, GCP and GVP training on every new or revised document and track completion for inspections, maintain qualification records for QPs, auditors and operators, administer onboarding, process time and attendance and answer policy questions. Hiring and appraisal decisions stay with people.
Systems validated, access reviewed, audit trails checked.
Agents maintain validation documentation for GxP systems, provision access and run periodic reviews, prepare audit-trail review packages, monitor the interfaces between ERP, LIMS, safety, RIM and serialization systems and triage security alerts, so IT keeps the validated state at scale.
Agents run inside your infrastructure, on-prem, private cloud or hybrid. No data leaves your network without your approval. No dependency on us: it runs on infrastructure you control. The agent is yours to run, permanently.
You pay for the process automated, not consulting hours. If a process proves non-automatable, a voucher covers the next one.
03
From 12 weeks · fastest go-live
Our fastest projects go live from 12 weeks; the plan for yours comes out of discovery, because it depends on the process and the systems it touches. Payback within one year for the large majority of projects.
Answers before the first call: the questions quality, regulatory and IT leaders ask us most.
How do you validate agents that touch GxP processes?
As any other computerized system under EU GMP Annex 11: a documented intended use, risk assessment, test evidence, an audit trail on every action and output, and change control on every update. The agent's outputs are records a person reviews and approves; the Qualified Person's release, the QPPV's causality assessment and medical judgment are never automated.
Do we need to replace our RIM, QMS, safety database or ERP?
No. Our agents sit on top of Veeva Vault, TrackWise, MasterControl, Argus, LifeSphere, SAP and your serialization platform and treat them as the systems of record they already are. Where an integration does not exist, agents operate through the same interfaces your staff use today.
Inside your environment: on-prem, private cloud or your own hybrid setup. Patient data from safety cases and clinical trials, unpublished dossiers and batch records stay in your environment by default, and access follows the same roles and information barriers you apply to people.
They measure technical automation potential: how much of the human work in a process is technically automatable at full deployment with clean, integrated data. They are not a forecast of headcount reduction and not a promise of realized savings. Laboratory and manufacturing execution and regulated decisions are not counted.
We do not automate them. Under the EU AI Act, screening candidates and evaluating performance are high risk and require human oversight. Agents track training and qualification records; hiring, appraisal and promotion decisions stay with your managers.
How do you price, and how long does it take?
Fixed price per workflow automation. If during delivery we determine a process cannot be automated, you receive a voucher to automate a different process instead. Our fastest projects go live from 12 weeks; the plan for yours comes out of discovery, and validated GxP deployments add the qualification steps your SOPs require.